The federal government is orchestrating a fundamental shift in how it regulates and evaluates psychedelic medicine, moving swiftly to bridge the gap between long-standing drug prohibitions and a surging national mental health emergency. Across federal agencies and executive leadership, a concerted effort is underway to compress review timelines, rethink clinical trial requirements, and prepare the nation’s healthcare infrastructure for what officials predict will be unprecedented public demand.
Writing recently in the New England Journal of Medicine, Food and Drug Administration officials unveiled an evolving framework specifically tailored to accelerate patient access to psychedelic compounds such as psilocybin and DMT. Acknowledging that existing psychiatric medications fail a substantial portion of the population, the agency emphasized that the worsening mental health crisis demands faster solutions. To compress the journey from laboratory bench to bedside, the agency is exploring ways to combine rolling reviews and priority review designations with conditional initial approvals. Under this model, treatments showing robust efficacy over a meaningful twelve-week window could reach patients while ongoing, long-term safety and dosing durability assessments continue through post-marketing studies and Risk Evaluation and Mitigation Strategies.
A central challenge the FDA aims to solve involves the methodological hurdles unique to psychedelic clinical trials, most notably the issue of functional unblinding. Because psychoactive compounds produce intense subjective experiences, trial participants and clinicians almost immediately recognize who received an active compound and who received an inactive placebo. While the agency still expects traditional control data to untangle drug-induced adverse events from baseline psychiatric conditions, officials have signaled a willingness to consider novel trial designs, including the use of alternative, active comparators that mimic altered states of consciousness rather than relying solely on inert placebos.
This administrative flexibility directly complements aggressive political momentum from the Department of Health and Human Services. HHS Secretary Robert F. Kennedy Jr. has repeatedly emphasized that federal regulators must mobilize rapidly, predicting an enormous surge in demand once these therapies receive legal clearance. Kennedy highlighted an executive order signed earlier this year by President Donald Trump designed to dismantle historical research roadblocks and streamline therapeutic pathways, crediting direct appeals from military veterans for catalyzing decisive action from the White House.
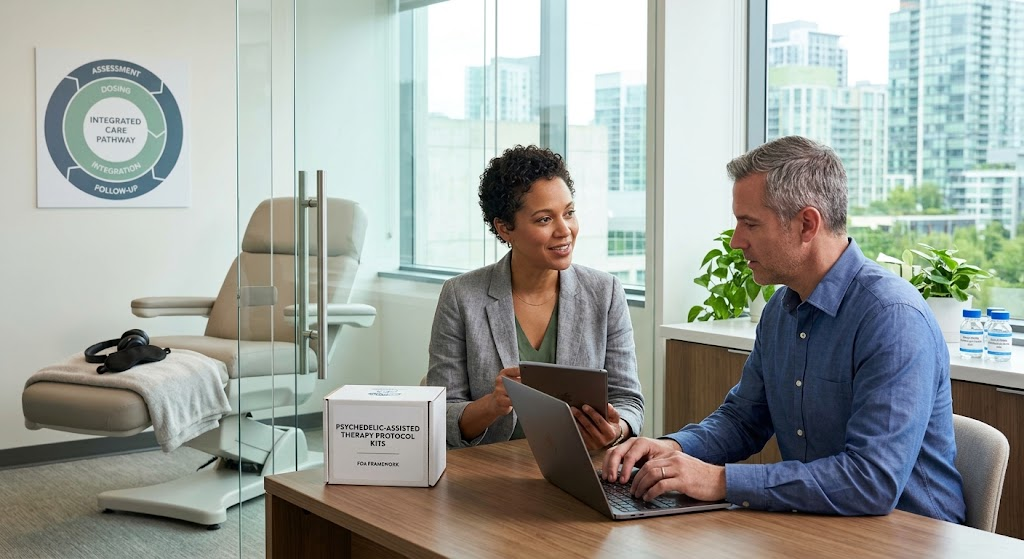
Yet, the transition from regulatory approval to practical care poses immense logistical and economic challenges. Kennedy pointed out that psychedelic-assisted therapy is far more resource-intensive than standard pharmacotherapy, requiring upwards of one hundred hours of trained clinical support staff per patient. While the Department of Veterans Affairs has explored mechanisms such as government-funded vouchers to help cover outside clinical care for veterans battling post-traumatic stress disorder and severe depression, standardizing these environments remains a daunting task.
Bridging that gap will require a massive transformation across both federal agencies and the medical establishment. As the FDA moves to establish gold-standard clinical protocols, standardize clinician training, and create post-market surveillance safeguards, the broader healthcare sector faces the reality that approving a compound is only half the battle. With bipartisan support in Congress and unprecedented coordination between the FDA, HHS, and the VA, the federal apparatus is now racing against the clock to ensure that when these breakthrough treatments finally arrive, the system is equipped to deliver them safely and ethically at scale.
Dabbin-Dad Newsroom